ZNG Medical
Medical device quality assurance is not a final inspection performed before shipment. It is a disciplined process that protects patients from design errors, unstable manufacturing, and unnoticed supplier changes. A reliable system begins with clear user needs, measurable specifications, and documented risk controls. Engineers must connect each requirement to verification evidence. They must also explain why the evidence is sufficient.
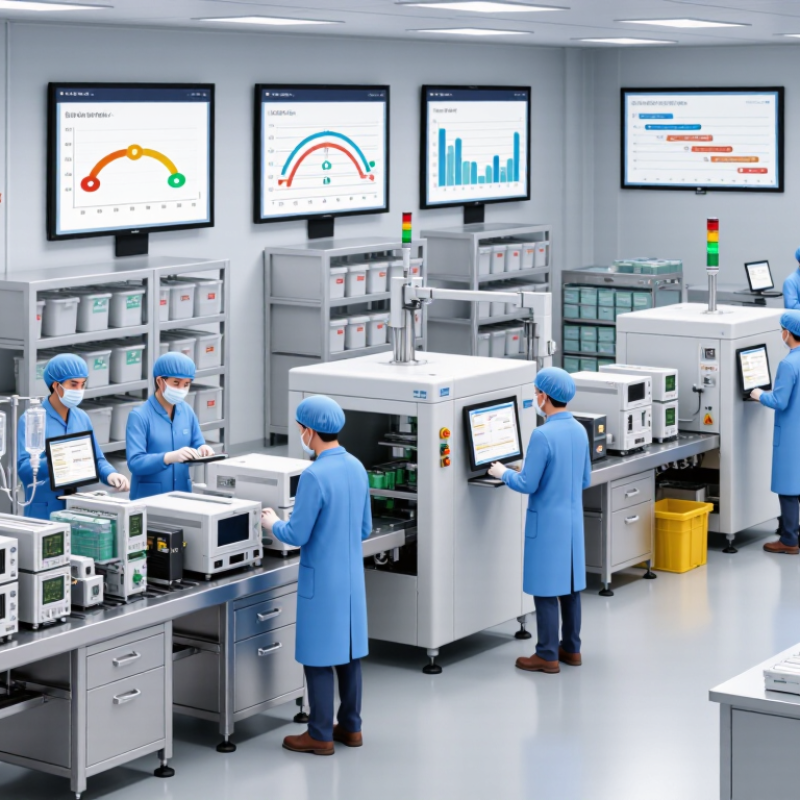
This guide presents practical Medical Device Quality Assurance Tips for teams developing, producing, or servicing medical devices. It examines design reviews, process validation, supplier qualification, equipment calibration, and production traceability. A well-controlled production line leaves visible clues: labeled components, approved work instructions, recorded temperatures, and complete batch histories. Complaint trends should reach quality teams quickly. Small signals can reveal serious weaknesses.
Standards such as ISO 13485 and ISO 14971 provide useful structure, but certificates alone cannot guarantee safe performance. Effective organizations test assumptions, investigate deviations, and verify corrective actions after implementation. CAPA records should identify root causes, not merely repeat training. Internal audits also need independence and practical judgment. No quality system is flawless. That admission matters. Teams can miss a weak seal, misread a supplier record, or accept incomplete validation data. Honest review creates room for correction. Strong medical device quality assurance combines technical knowledge, frontline experience, regulatory awareness, and consistent documentation. It turns quality from paperwork into dependable patient protection.
Medical device quality assurance is the planned, documented work that helps ensure a device performs safely and consistently throughout its life cycle. It is broader than inspecting finished products. Teams set requirements, control suppliers, validate manufacturing processes, review complaints, and track corrective actions. The World Health Organization’s Global Atlas of Medical Devices reports roughly 2 million types of devices, grouped into about 7,000 generic categories. That scale makes clear why one checklist cannot fit every device. A sterile dressing and a diagnostic instrument need different controls.
Core objectives are patient safety, reliable performance, and traceable evidence. In practice, that means linking a design requirement to a test result, a production record, and a response when something fails. FDA’s 2024 Quality Management System Regulation final rule aligns U.S. requirements more closely with ISO 13485, reinforcing a lifecycle approach to quality systems. Good records matter. A missing calibration entry can undermine confidence in otherwise sound measurements. Quality assurance also uses complaint trends and audit findings to identify risks before they recur. Yet procedures can look complete while staff struggle to follow them on a busy production line. That gap deserves honest review: observe the work, ask operators where steps break down, and revise controls without weakening safety.
| Core Objective | What It Means | Typical Quality Controls | Examples of Objective Evidence | Useful Monitoring Indicators |
|---|---|---|---|---|
| Define and maintain a quality management system | Establish documented processes that consistently meet applicable regulatory and product requirements throughout the device life cycle. | Assign responsibilities; control procedures and records; conduct internal audits and management reviews; address nonconformities. | Approved procedures, training records, audit reports, management-review records, and corrective-action records. | Audit findings by category; overdue corrective actions; recurring nonconformities. |
| Design safe and effective devices | Translate user needs and intended use into verified design inputs and validated design outputs, while managing design changes. | Design planning and reviews; requirements traceability; verification; validation; controlled design transfer and change review. | Design plans, review minutes, test reports, validation records, traceability matrices, and change approvals. | Unresolved design issues; verification and validation completion; changes requiring additional assessment. |
| Identify and control risk | Identify hazards, estimate and evaluate risks, implement risk controls, and review residual risk using a documented process. | Risk analysis across the life cycle; implementation and verification of controls; review of production and post-production information. | Risk-management records, hazard analyses, control verification results, and documented residual-risk evaluations. | Open risk-control actions; risk-related complaints; trends in reported incidents or safety signals. |
| Ensure consistent supplier and production quality | Make sure externally supplied products and production processes meet defined requirements and are controlled proportionately to risk. | Supplier evaluation and monitoring; purchasing controls; validated processes where output cannot be fully verified later; production monitoring. | Supplier assessments, purchase specifications, incoming inspection records, process-validation records, and production-device history records. | Supplier nonconformities; incoming acceptance results; process deviations; production yield and rework trends. |
| Verify product conformity before release | Confirm that each applicable product or batch meets specified acceptance criteria before it is released for distribution. | Inspection and testing; calibrated measuring equipment; review of required production records; authorized release. | Inspection results, test records, equipment-calibration records, completed production records, and release authorization. | First-pass acceptance; rejected or reworked units; release-record completion and review time. |
| Maintain traceability and control nonconforming product | Identify product status, trace product where required, and prevent unintended use or distribution of nonconforming product. | Identification and status controls; lot, batch, or serial-number records as appropriate; segregation and documented disposition. | Traceability records, nonconformance reports, investigation records, disposition approvals, and rework verification. | Time to identify affected product; open nonconformance age; recurrence of similar defects. |
| Use post-market information to improve quality | Collect and evaluate relevant field information, respond to feedback and complaints, and take appropriate corrective or preventive action. | Complaint handling; investigation and trend analysis; applicable reporting processes; corrective and preventive action; effectiveness checks. | Complaint files, trend reports, investigation records, field-action documentation where applicable, and effectiveness-check records. | Complaint trends by issue type; investigation timeliness; corrective-action effectiveness; repeat issues after closure. |
Definition: Medical device quality assurance is the planned, systematic set of activities used to provide confidence that a medical device and its supporting processes consistently meet applicable requirements. Indicators should be interpreted against documented objectives, risk, device type, and applicable regulations; no single target is appropriate for every device.
It covers planned, documented controls throughout a device’s life cycle. These include design, suppliers, manufacturing, complaints, and corrective actions. Not just final inspection.
The core goals are patient safety, reliable performance, and traceable evidence. A design requirement should connect to a test, a production record, and a response when something fails.
Identify hazards, estimate their risks, and link each control to a requirement and test. Rank risks by severity and likelihood. Keep those links visible in a traceability matrix.
Test plausible failures, not only normal operation. A temperature monitor might need checks for sensor drift, low-battery behavior, and alarm visibility in bright light. One passing test is not enough.
Reports can reveal patterns for further investigation. They do not establish event rates or prove that a device caused an event. Treat them as signals, not conclusions.
Track measures such as seal strength, molding dimensions, and sterilization-cycle settings. Review results by machine, shift, and material lot. Averages can hide small changes.
Check material specifications, change notifications, and traceability records. Compare supplier certificates with incoming inspection results. Annual questionnaires may miss practical problems.
Keep records legible, dated, and linked to specific lots. Document calibration changes, reasons, and approvals. Verify corrective actions work in practice, not just on paper. A checklist can still miss things.
Medical Device Quality Assurance is a structured approach to ensuring that a device is safe, reliable, and performs as intended throughout its lifecycle. Effective quality planning begins by defining clear objectives, assigning responsibilities, and mapping applicable regulatory requirements to the organization’s quality management processes. Medical Device Quality Assurance Tips include integrating risk assessment into design decisions, setting measurable acceptance criteria, and using planned testing and verification to identify issues before production.
Quality assurance also depends on consistent manufacturing controls, qualified suppliers, accurate records, and regular process monitoring. These practices help teams detect variation, investigate problems, and document corrective actions. After a device reaches the market, organizations should review performance data, user feedback, and reported quality concerns to spot emerging trends. Using these findings to improve designs, procedures, and training supports continuous improvement and helps maintain product quality over time.